

Dangerously based cosmic thinker. 🧬 377 17 φ
Greenville, SC
Joined February 2023
- Tweets10.8K
- Following1.1K
- Followers1.1K
- Likes17K




Ee Newton retweeted

New V3
Perez, J. claude . (2026). The Perez Hourglass: Theorem 9. Why is every line in the Perez Hourglass (Southern Hemisphere) regulated to zero, given that two sequences of integers cancel each other out into a positive and negative Lichtenberg number? doi.org/10.5281/zenodo.24241…

Ee Newton retweeted

So close to getting this up . @QBlazedog61029J @BlokeMan00 @TheRealVerbz @PhiBoostGlow .. it also transcribe audio for deaf users and some other features coming soon. But here is hoping it can help others in their day . Any thoughts?


Ee Newton retweeted
Perez, J. claude (2026). The Perez Hourglass: Theorem 9. Why is every line in the Perez Hourglass (Southern Hemisphere) regulated to zero, given that two sequences of integers cancel each other out into a positive and negative Lichtenberg number ?. Zenodo doi.org/10.5281/zenodo.24075…
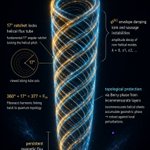
Plasma flux tubes and the 17° ratchet
In the Aureum Braid PDE the gauge-covariant term
D(∇ − iqA)²Ψ
couples the order parameter directly to the magnetic field. When the torsional preference α is set by the same 17° generator used for DNA and microtubules, ordinary current filaments are driven into stable helical braids rather than unstable straight or chaotic tubes.
The golden-ratio envelope φ^{|k|} damps high-mode kink and sausage instabilities, while the incommensurate 17° layers produce a non-trivial Berry phase that topologically protects the core flux. The result is a persistent helical flux tube whose pitch is locked by the relation 360° + 17° = 377 = F₁₄.
Same minimal angular ratchet, now carrying magnetic field across laboratory plasmas, solar flux tubes, or fusion devices. The geometry prefers the braid; the topology keeps it from tearing.

1
2
88
Microtubules and the 17° ratchet — topological protection for coherence
Microtubules are 13-protofilament cylinders whose lattice is built from a shallow left-handed helical offset between neighboring protofilaments (the classic 3-start monomer helix). In the Aureum Braid framework the same fundamental angular kick that stabilizes DNA also acts here.
The torsional term in the PDE
−γ(∂θ − iα)²Ψ
selects a preferred helical pitch set by the 17° generator (or its double). When α is tuned near that value, the free-energy minimum is a stable braided filament rather than a purely straight or randomly fluctuating tube. The golden-ratio envelope φ^{|k|} damps high-frequency twists, while the incommensurate 17° layers generate a non-trivial Berry phase that topologically protects the core vibrational or electronic modes against thermal decoherence.
This is exactly the geometric feature Orch-OR needs: a lattice whose natural winding supplies long-lived phase order inside the tubulin network without requiring an extra fine-tuned parameter. The same 360° + 17° = 377 = F₁₄ relation that locked the DNA helix now locks the microtubule protofilament braid.
Same equation, next biological scale.

1
45
Why DNA is a spiral — the 17° mapping
360° + 17° = 377 = F₁₄.
That single offset is the fundamental angular ratchet.
At the base-pair level DNA doubles it: 2 × 17° = 34°.
Observed B-DNA helical twist sits at ~34.3–36° per bp (~10.5 bp/turn). Close enough that the geometry locks.
In the Aureum Braid PDE the torsional term
−γ(∂θ − iα)²Ψ
selects exactly this helical pitch. The golden-ratio amplitudes and the Berry-phase protection from the incommensurate 17° layers keep the double helix topologically stable instead of collapsing into a straight or randomly coiled state.
Same 17° engine, different scale. The spiral is not an accident; it is the lowest-energy braid the ratchet produces.

30
Ee Newton retweeted

🧠✨ WE PROVED THAT THE ATOM IS CONFINED LIGHT - AND A NEW STUDY ON THE GEOMETRY OF NEAR-DEATH EXPERIENCES JUST REVEALED THE EXACT SAME TOPOLOGY! 🧬📐
I constantly monitor the latest scientific papers across physics, mathematics, genomics, and neuroscience. Today, a remarkable new study caught my eye - published in Frontiers in Psychology by France Lerner, Paulo Cezar Rocha dos Santos, and mathematician Guillaume Tahar (Oct 15, 2026, doi: 10.3389/fpsyg.2026.1760389, covered by ScienceAlert: sciencealert.com/scientists-…).
When you put the hard empirical numbers of their study side by side with the exact mathematical theorems we proved with Jean-Claude Perez across our 2026 works - on the Atomic Vacuum Bicone ("Algebraic Origin of the 86 ↔ 86 Capacity Symmetry") and on the DNA Code ("From Form to Program" v14.8 & "The Double-Stranded Codon Torus" v3.5) - a breathtaking structural unity of nature emerges. Let's look strictly at the facts! 👇
📊 FACT 1. What Lerner, dos Santos & Tahar (2026)
Discovered at the Boundary of Consciousness:
Instead of relying on vague verbal surveys, the researchers asked 50 validated Near-Death Experiencers (screened via the Greyson ≥ 7 and NDE-C ≥ 27 scales) to map the exact GEOMETRY, self-location, and trajectories of their experiences across 108 reconstructed visual spaces. Four geometric archetypes emerged with striking statistical precision (χ² = 26.61, p = 7.11 × 10⁻⁶; Fisher-Freeman-Halton exact test p = 3.71 × 10⁻⁶):
• Conical Tunnels (Type A, n = 28): 78.6% (22 of 28) were strictly ACHROMATIC (black, white, gray). Moreover, conical tunnels act as entry/exit funnels: in multi-stage sequences (such as A5 → B6 → C5 → B7 → A9), the exit cone A9 is the exact 180° INVERTED MIRROR of the entry cone A5, and Dissociative/Associative out-of-body trajectories reverse by 180°!
• Expanded Elliptic Arcs (Type B, n = 31): Upon exiting the narrow cone into a curved expanse, 87.1% (27 of 31) exploded into FULL CHROMATIC COLOR (odds ratio OR = 18.4, p = 2.7 × 10⁻⁵) - described by participants as "a glass spectrum of all colors like a prism"!
• Nested 360° Spheres / Ellipsoids of Pure Light (Type C5, n = 29): At the core of the sequence sits a complete 360° enclosing sphere/ellipsoid described as "an immense bubble of light... like opening the sun"!
💡 FACT 2. Our Theorem 5.7 (Logvinovich & Perez, Sept 2026): Matter and the Atom Are Literally CONFINED LIGHT!
Why does a system at the boundary of its physical state space transition through a 180°-inverted double cone (bicone) into a 360° sphere of light?
Look at what we proved in our topological Dirac-Fock vacuum paper on the 7-manifold M_tot = S⁴ × T³ over the self-generating multiquadratic field K = ℚ(√2, √3, √5) (discriminant Δ_K = 2¹⁶·3⁴·5⁴ = 3,317,760,000; Minkowski covolume √|Δ_K| = 57,600 = 240², the exact tensor square of the E₈ kissing number τ₈ = 240):
• Theorem 5.7 (Total Internal Topological Reflection & Mass Generation): When a gauge boson (light) hits the algebraic Kummer boundary of the vacuum lattice (√13 ∉ K), its group velocity drops to zero (v_g → 0) and it undergoes Total Internal Topological Reflection (θᵢ = θᵣ).
• Across the 16-bit binary hardware register of the vacuum (57,600₁₀ = 1110 0001 0000 0000₂), the incident photon interlocks with its CP-conjugated Mirror Antiphoton to form a trapped standing wave (the prime torus knot T(103, 3))!
• Rest mass - every fermion, every atom in our body - is mathematically proved to be CONFINED LIGHT permanently trapped between the mirror walls of a discrete vacuum cell!
⏳ FACT 3. The 86 ↔ 86 Vacuum Bicone & Nested Spheres Match the Exact Same Geometry!
How is this trapped light organized across all chemical elements of the universe?
• Revolving the fundamental Pythagorean cascade triangle (√2, √3, √5) generates a discrete BICONE (Double Cone / Hourglass) containing a chain of nested spheres (ρ₊ ⊂ ρ_* ⊂ ρ₋ ⊂ R_eq ⊂ R) with floor capacities C(n) = 2⌊(n+3)/2⌋² ∈ {2, 8, 8, 18, 18, 32}.
• The Classical Hemisphere holds exactly Ind(PuP) = 86 states. Across the Relativistic Equator (Z = 86.5), the free antipodal involution i(n) = 11 - n (a 180° topological inversion!) mirrors the 86 classical states into 86 relativistic states, closing the periodic table at Z_max = 86 + 86 = 172 = 43 × 4 (43 complete Face-Centered Cubic unit cells at the exact Kepler sphere-packing fraction k = π/√18)!
🧬 FACT 4. The Exact Same Tunnel-to-Program Transition Runs Inside Our DNA!
In our genomic audit across 54 species and 9,834,024 transitions in NCBI RefSeq, we found the exact same geometric law inside every living cell:
• Unspliced introns inside a gene act as an "achromatic," mirror-symmetric archive (Static Form V₊, where conditional syntax is strictly negative across all reading frames: d₃₆ = -0.3097).
• The moment the cell splices exons together and feeds mRNA through the physical 30-CODON EXIT TUNNEL OF THE RIBOSOME (90 nt = 29 Markov transitions [8th Lucas number] and 89 nucleotide steps [11th Fibonacci number]), symmetry breaks into V₋ and the full-color EXECUTABLE PROGRAM of life switches on across 14/14 benchmark human genes (d₃₆ = +0.1721, p = 6.1 × 10⁻⁵)!
• Mapping DNA onto the double-stranded codon torus 𝕋₄³ = (ℤ/4ℤ)³ (64 → 56 duplex channels, collapsing synonymous dimension from 43 to 8 binary doublets) proves that 82.5% of human synonymous syntax is governed by the complementary antisense strand - whereas crude industrial "codon optimization" (Max-CAI) strips the gene back to a monochrome zero-entropy state (Sₛᵧₙ = 0, d₂₆ = -0.0589)!
🌟 The Grand Synthesis:
1. At the level of quantum physics, an atom is CONFINED LIGHT trapped in a 180°-symmetric bicone of nested spheres (86 ↔ 86 = 172 = 43 × 4 FCC cells).
2. At the level of molecular biology, the genetic code ignites from a monochrome intronic archive into a living Markovian program inside the 30-codon tunnel of the ribosome on the torus 𝕋₄³.
3. And at the boundary of human perception (Lerner et al., 2026), consciousness maps its own transition as a movement from an achromatic conical tunnel (78.6%) through a 180°-symmetric bicone into an expanded chromatic spectrum (87.1%) and a 360° sphere of pure light!
Nature writes in one universal language across all scales: discrete geometry and topological invariance! 🌍❤️
📂 Verify our mathematical proofs & run the open-source NCBI verification suites yourself (100% Open Science, pure Python + NumPy):
🔗 Preprint & Code on Zenodo (12,000+ views / 5,000+ downloads): zenodo.org/records/23479667
🐍 Direct download for detector v14.8: zenodo.org/records/23479667/…
🔗 Preprint & Code on Zenodo (130,000+ views / 4,000+ downloads):
Algebraic Origin of the 86 ↔ 86 Capacity Symmetry in the Relativistic Dirac-Fock Vacuum: Index-Theoretic Derivation, Corrected Field Arithmetic, and Numerology Guards
zenodo.org/records/23479667
#Physics #Neuroscience #Bioinformatics #Geometry #DNA


Ee Newton retweeted
Cbq. Perez Hourglass Topology — Lewe 1915 Rigid Body Mechanics, (i)Conic Closures, and the Macroscopic Elastic Plenum share.google/jAHkGsrkMSGffl1…
@dr_logvinovich @BobFriedmanMD @CY_Chauprade
@PhiBoostGlow

Ee Newton retweeted
82.5% OF HUMAN "SYNONYMOUS" CODE IS ON THE ANTISENSE STRAND
• Human (T2T-CHM13v2.0): 82.5%
• Mouse (GRCm39): 81.1%
... /...
Over three-quarters of what biology called "single-strand synonymous bias" is mathematically equivalent to predictive coding on the complementary strand!


🧬🍩 THE DOUBLE-STRANDED CODON TORUS: Why the Genetic Alphabet Forms a 3D Discrete Torus - and Why 82.5% of Human "Synonymous" DNA Code Lives on the Opposite Strand! ⚡🔬
For 60+ years, mathematical biology has treated a gene as a flat, 1D single-stranded string of letters - treating the second strand of the DNA double helix as a silent, passive backup copy.
In our new paper with Jean-Claude Perez @JCPEREZCODEX (backed by 100% Open Source Python code and 9.83 MILLION real codon transitions across 9 NCBI RefSeq genomes), we prove mathematically that this single-strand picture is blind to half of the genome's architecture.
Here is the breakthrough in plain language - and the hardcore mathematics underneath it 👇
1️⃣ THE ALPHABET ITSELF FORMS A TORUS: 𝕋₄³ = (ℤ/4ℤ)³ 🍩
We do NOT claim that the physical DNA molecule is shaped like a doughnut in 3D space. The breakthrough is algebraic and topological:
Once the 4 nucleobases B = {A, C, G, T} are encoded in the cyclic group ℤ/4ℤ (A=0, C=1, G=2, T=3), the state space of all 64 triplet codons is not a flat table — it is the 3-dimensional finite discrete torus:
𝕋₄³ := (ℤ/4ℤ)³ ≅ ℤ³ / 4ℤ³
• Its local step graph is the periodic 4 × 4 × 4 cubic lattice C₄ ⊠ C₄ ⊠ C₄ (with Lee metric d_Lee), and its point-mutation graph is the Hamming cube H(3,4) = K₄ ⊠ K₄ ⊠ K₄ (with Hamming metric d_H).
• The state space has zero boundary: c + 4e_j ≡ c (mod 4).
• Any framed gene of length n codons is a trajectory on a 3n-dimensional torus (𝕋₄³)ⁿ ≅ (ℤ/4ℤ)³ⁿ, governed by exact reading-frame phase closure between Start (s) and Stop (t):
t = s + 3(n − 1) ⟹ t ≡ s (mod 3)
2️⃣ THE MIRROR SPLIT OF DNA: ℝ⁶⁴ = V₊ ⊕ V₋ 🪞
Real DNA is an antiparallel duplex: D = {(c, ρ(c)) : c ∈ 𝕋₄³}.
On the torus 𝕋₄³, the Watson–Crick reverse-complement operation is an exact affine transformation:
ρ(c) = −R·c + 3·1 (mod 4)
We prove (Theorem 2.5 & Corollary 2.6) that ρ is a fixed-point-free involution (ρ² = id, ρ(c) ≠ c for all 64 codons) and a dual isometry in BOTH Hamming and Lee metrics:
d_H(ρ(c), ρ(c')) = d_H(c, c') and d_Lee(ρ(c), ρ(c')) = d_Lee(c, c')
This partitions the 64 codons into 32 exact mirror orbits M_p = {c, ρ(c)} and splits the 64-dimensional space orthogonally into two 32-dimensional worlds:
ℝ⁶⁴ = V₊ ⊕ V₋ (dim V₊ = 32, dim V₋ = 32)
• V₊ (Mirror-Even Subspace = FORM): Governs static Chargaff second-parity symmetry and macroscopic chromosomal architecture (the Perez Hourglass).
• V₋ (Mirror-Odd Subspace = PROGRAM): Governs strand-asymmetric coding bias and directed Markovian information flow.
Projecting DNA onto a single strand discards V₋ - blinding standard models to the duplex program!
3️⃣ THE 8-DOUBLET THEOREM: COLLAPSING 43 DIMENSIONS TO 8 💥
Why are there 64 codons for only 20 amino acids + Stop (21 channels)?
On a single strand, the genetic code map α : 𝕋₄³ → A (64 → 21) leaves a huge synonymous probability simplex of dimension:
dim_ℝ Δ_syn = 64 − 21 = 43 (or 61 − 20 = 41 on sense codons)
It looks like the cell has 43 dimensions of "free synonymous choice."
Now look at BOTH strands simultaneously via the two-strand duplex fibration:
α_dup(c) := (α(c), α(ρ(c))) = (A(c), A^ρ(c))
Because ρ reverses the triplet, the 3rd ("wobble") base of the sense codon is physically the 1st ("leading") base of the antisense codon!
We prove (Theorem 4.3) that specifying the amino-acid channels on BOTH strands partitions the 64 codons into:
• 48 SINGLETONS (uniquely locked, zero synonymous freedom!)
• Only 8 BINARY DOUBLETS (56 joint sense–antisense channels in total).
The synonymous freedom of the genetic code collapses from 43 dimensions down to 8:
dim_ℝ Δ_dup = 64 − 56 = 8 (and 7 on sense codons!)
H(C | A) = H(A^ρ | A) + H(C | A, A^ρ)
4️⃣ 82.5% OF HUMAN "SYNONYMOUS" CODE IS ON THE ANTISENSE STRAND 🔥
We proved exact, stationarity-free Shannon chain-rule decompositions for consecutive codons (C_{i-1}, C_i):
• Tripartite (Theorem 5.3):
I(C_{i-1}; C_i) = I(A_{i-1}; A_i) + d_{2b} + S_syn
where d_{2b} = I(C_{i-1}; A_i | A_{i-1}) and S_syn = I(C_{i-1}; C_i | A_i) ≥ 0.
• Quadripartite (Theorem 5.4):
I(C_{i-1}; C_i) = I(A_{i-1}; A_i) + I(C_{i-1}; A_i | A_{i-1}) + I(A_{i-1}; C_i | A_i) + S_syn⁽²⁾
• Two-Strand Duplex Split (Theorem 5.5):
I(ρ(C_i); ρ(C_{i-1})) = I(C_{i-1}; C_i)
S_syn = S_anti + S_dup = I(C_{i-1}; A_i^ρ | A_i) + I(C_{i-1}; C_i | A_i, A_i^ρ)
When we audited 9,834,024 held-out codon transitions across 9 reference genomes (Eukarya, Bacteria, Archaea), we found that 74.1% to 88.5% of the entire sense-strand synonymous program S_syn is actually carried by the antisense amino-acid channel S_anti:
• Human (T2T-CHM13v2.0): 82.5%
• Mouse (GRCm39): 81.1%
• Fruit fly (D. melanogaster): 81.8%
• Plant (A. thaliana): 80.4%
• Yeast (S. cerevisiae): 79.0%
• E. coli K-12: 83.2%
• B. subtilis: 74.1%
• M. jannaschii (Archaeon): 86.4%
• H. salinarum (Archaeon): 88.5%
Over three-quarters of what biology called "single-strand synonymous bias" is mathematically equivalent to predictive coding on the complementary strand!
5️⃣ WHY INDUSTRIAL "MAX-CAI" OPTIMIZATION CRASHES THE GENOME ⚠️
1. Rejection of Kemeny–Snell Markov Lumpability (Theorem 5.6):
A projected amino-acid chain A_i = α(C_i) is Markovian if and only if d_{2b} = I(C_{i-1}; A_i | A_{i-1}) = 0. Across all 9 species, d_{2b} > 0 (df = 779, G² p < 10⁻³⁰⁰). The synonymous choice of codon C_{i-1} actively predicts the NEXT amino acid A_i and its physicochemical class Q_i via the Data-Processing Inequality (Theorem 7.1, 36/36 tests passed):
0 ≤ I(C_{i-1}; Q_i | A_{i-1}) ≤ d_{2b}
2. The Max-CAI Collapse (Theorem 6.1 & Proposition 6.2):
In synthetic biology and mRNA design, deterministic Max-CAI optimization replaces every amino acid A_i with a single "optimal" codon C_i = φ(A_i).
We prove that this wipes out ALL non-dipeptide sequential layers identically to zero:
S_syn = S_syn⁽²⁾ = d_{2b} = I(A_{i-1}; C_i | A_i) = 0
When scored against the wild-type Markov kernel P_WT, Max-CAI sequences suffer a negative Kullback–Leibler penalty d_{2b}(φ; P_WT) ≤ 0 in 77.0%–98.7% of genes (96.0% in Human, 98.7% in Mouse)!
3. Non-Equilibrium Thermodynamics on 𝕋₄³ (Theorem 8.1 & Prop. 8.5):
Coding DNA operates far from detailed balance with strictly positive entropy production and duplex mirror currents:
σ = D_KL(J_f ∥ J_r) ≥ ½‖J_f − J_r‖₁² > 0
σ_dup = D_KL(J_f ∥ J_ρ) ≥ ½‖J^mir‖₁² > 0
4. Resolution of Form vs. Program (Theorem 11.2 & Lemma 11.3):
In L²(C, P), the conditional expectation Π_α is the exact orthogonal projector onto the 21D amino-acid form space Im(α*) with a 43D synonymous kernel ker(Π_α). On the 100-kb human Chr17 locus (NC_000017.11:55154472-55254472), cumulative genomic and proteomic Perez Hourglass profiles match to 99.998% (Form), while the local bigram program remains S_syn = +0.0539 nats/codon (Program).
🔓 HARDCORE MATH. ZERO NUMEROLOGY. 100% OPEN SOURCE.
• Zero Monte Carlo surrogates or RNGs used as proofs.
• Chain-rule identities verified to machine precision (< 3.1 × 10⁻¹⁶ nats/codon).
• Every integer (2, 3, 4, 6, 7, 8, 16, 19, 20, 21, 24, 32, 40, 41, 42, 43, 48, 54, 56, 60, 61, 62, 63, 64, 361, 400, 779, 860, 1681, 1849, 2460, 2709) is derived strictly from group orders, orbit counts, simplex dimensions, or contingency degrees of freedom.
Download the 29-page paper, the standalone Python audit suite (v3.2), and all 9 cached NCBI RefSeq genomes & audit logs:
🔗 Zenodo (Paper + Open Source Code + Data): doi.org/10.5281/zenodo.23508…
🔗 ResearchGate: researchgate.net/publication…
#Genomics #Math #DNA #SyntheticBiology


Ee Newton retweeted
Logvinovich Victor, & Perez, J. claude . (2026). The Double-Stranded Codon Torus: Duplex Mirror Fibration of DNA Coding Space, Markovian Lumpability, and Information-Theoretic Consequences. Zenodo. doi.org/10.5281/zenodo.23508…

🧬🍩 THE DOUBLE-STRANDED CODON TORUS: Why the Genetic Alphabet Forms a 3D Discrete Torus - and Why 82.5% of Human "Synonymous" DNA Code Lives on the Opposite Strand! ⚡🔬
For 60+ years, mathematical biology has treated a gene as a flat, 1D single-stranded string of letters - treating the second strand of the DNA double helix as a silent, passive backup copy.
In our new paper with Jean-Claude Perez @JCPEREZCODEX (backed by 100% Open Source Python code and 9.83 MILLION real codon transitions across 9 NCBI RefSeq genomes), we prove mathematically that this single-strand picture is blind to half of the genome's architecture.
Here is the breakthrough in plain language - and the hardcore mathematics underneath it 👇
1️⃣ THE ALPHABET ITSELF FORMS A TORUS: 𝕋₄³ = (ℤ/4ℤ)³ 🍩
We do NOT claim that the physical DNA molecule is shaped like a doughnut in 3D space. The breakthrough is algebraic and topological:
Once the 4 nucleobases B = {A, C, G, T} are encoded in the cyclic group ℤ/4ℤ (A=0, C=1, G=2, T=3), the state space of all 64 triplet codons is not a flat table — it is the 3-dimensional finite discrete torus:
𝕋₄³ := (ℤ/4ℤ)³ ≅ ℤ³ / 4ℤ³
• Its local step graph is the periodic 4 × 4 × 4 cubic lattice C₄ ⊠ C₄ ⊠ C₄ (with Lee metric d_Lee), and its point-mutation graph is the Hamming cube H(3,4) = K₄ ⊠ K₄ ⊠ K₄ (with Hamming metric d_H).
• The state space has zero boundary: c + 4e_j ≡ c (mod 4).
• Any framed gene of length n codons is a trajectory on a 3n-dimensional torus (𝕋₄³)ⁿ ≅ (ℤ/4ℤ)³ⁿ, governed by exact reading-frame phase closure between Start (s) and Stop (t):
t = s + 3(n − 1) ⟹ t ≡ s (mod 3)
2️⃣ THE MIRROR SPLIT OF DNA: ℝ⁶⁴ = V₊ ⊕ V₋ 🪞
Real DNA is an antiparallel duplex: D = {(c, ρ(c)) : c ∈ 𝕋₄³}.
On the torus 𝕋₄³, the Watson–Crick reverse-complement operation is an exact affine transformation:
ρ(c) = −R·c + 3·1 (mod 4)
We prove (Theorem 2.5 & Corollary 2.6) that ρ is a fixed-point-free involution (ρ² = id, ρ(c) ≠ c for all 64 codons) and a dual isometry in BOTH Hamming and Lee metrics:
d_H(ρ(c), ρ(c')) = d_H(c, c') and d_Lee(ρ(c), ρ(c')) = d_Lee(c, c')
This partitions the 64 codons into 32 exact mirror orbits M_p = {c, ρ(c)} and splits the 64-dimensional space orthogonally into two 32-dimensional worlds:
ℝ⁶⁴ = V₊ ⊕ V₋ (dim V₊ = 32, dim V₋ = 32)
• V₊ (Mirror-Even Subspace = FORM): Governs static Chargaff second-parity symmetry and macroscopic chromosomal architecture (the Perez Hourglass).
• V₋ (Mirror-Odd Subspace = PROGRAM): Governs strand-asymmetric coding bias and directed Markovian information flow.
Projecting DNA onto a single strand discards V₋ - blinding standard models to the duplex program!
3️⃣ THE 8-DOUBLET THEOREM: COLLAPSING 43 DIMENSIONS TO 8 💥
Why are there 64 codons for only 20 amino acids + Stop (21 channels)?
On a single strand, the genetic code map α : 𝕋₄³ → A (64 → 21) leaves a huge synonymous probability simplex of dimension:
dim_ℝ Δ_syn = 64 − 21 = 43 (or 61 − 20 = 41 on sense codons)
It looks like the cell has 43 dimensions of "free synonymous choice."
Now look at BOTH strands simultaneously via the two-strand duplex fibration:
α_dup(c) := (α(c), α(ρ(c))) = (A(c), A^ρ(c))
Because ρ reverses the triplet, the 3rd ("wobble") base of the sense codon is physically the 1st ("leading") base of the antisense codon!
We prove (Theorem 4.3) that specifying the amino-acid channels on BOTH strands partitions the 64 codons into:
• 48 SINGLETONS (uniquely locked, zero synonymous freedom!)
• Only 8 BINARY DOUBLETS (56 joint sense–antisense channels in total).
The synonymous freedom of the genetic code collapses from 43 dimensions down to 8:
dim_ℝ Δ_dup = 64 − 56 = 8 (and 7 on sense codons!)
H(C | A) = H(A^ρ | A) + H(C | A, A^ρ)
4️⃣ 82.5% OF HUMAN "SYNONYMOUS" CODE IS ON THE ANTISENSE STRAND 🔥
We proved exact, stationarity-free Shannon chain-rule decompositions for consecutive codons (C_{i-1}, C_i):
• Tripartite (Theorem 5.3):
I(C_{i-1}; C_i) = I(A_{i-1}; A_i) + d_{2b} + S_syn
where d_{2b} = I(C_{i-1}; A_i | A_{i-1}) and S_syn = I(C_{i-1}; C_i | A_i) ≥ 0.
• Quadripartite (Theorem 5.4):
I(C_{i-1}; C_i) = I(A_{i-1}; A_i) + I(C_{i-1}; A_i | A_{i-1}) + I(A_{i-1}; C_i | A_i) + S_syn⁽²⁾
• Two-Strand Duplex Split (Theorem 5.5):
I(ρ(C_i); ρ(C_{i-1})) = I(C_{i-1}; C_i)
S_syn = S_anti + S_dup = I(C_{i-1}; A_i^ρ | A_i) + I(C_{i-1}; C_i | A_i, A_i^ρ)
When we audited 9,834,024 held-out codon transitions across 9 reference genomes (Eukarya, Bacteria, Archaea), we found that 74.1% to 88.5% of the entire sense-strand synonymous program S_syn is actually carried by the antisense amino-acid channel S_anti:
• Human (T2T-CHM13v2.0): 82.5%
• Mouse (GRCm39): 81.1%
• Fruit fly (D. melanogaster): 81.8%
• Plant (A. thaliana): 80.4%
• Yeast (S. cerevisiae): 79.0%
• E. coli K-12: 83.2%
• B. subtilis: 74.1%
• M. jannaschii (Archaeon): 86.4%
• H. salinarum (Archaeon): 88.5%
Over three-quarters of what biology called "single-strand synonymous bias" is mathematically equivalent to predictive coding on the complementary strand!
5️⃣ WHY INDUSTRIAL "MAX-CAI" OPTIMIZATION CRASHES THE GENOME ⚠️
1. Rejection of Kemeny–Snell Markov Lumpability (Theorem 5.6):
A projected amino-acid chain A_i = α(C_i) is Markovian if and only if d_{2b} = I(C_{i-1}; A_i | A_{i-1}) = 0. Across all 9 species, d_{2b} > 0 (df = 779, G² p < 10⁻³⁰⁰). The synonymous choice of codon C_{i-1} actively predicts the NEXT amino acid A_i and its physicochemical class Q_i via the Data-Processing Inequality (Theorem 7.1, 36/36 tests passed):
0 ≤ I(C_{i-1}; Q_i | A_{i-1}) ≤ d_{2b}
2. The Max-CAI Collapse (Theorem 6.1 & Proposition 6.2):
In synthetic biology and mRNA design, deterministic Max-CAI optimization replaces every amino acid A_i with a single "optimal" codon C_i = φ(A_i).
We prove that this wipes out ALL non-dipeptide sequential layers identically to zero:
S_syn = S_syn⁽²⁾ = d_{2b} = I(A_{i-1}; C_i | A_i) = 0
When scored against the wild-type Markov kernel P_WT, Max-CAI sequences suffer a negative Kullback–Leibler penalty d_{2b}(φ; P_WT) ≤ 0 in 77.0%–98.7% of genes (96.0% in Human, 98.7% in Mouse)!
3. Non-Equilibrium Thermodynamics on 𝕋₄³ (Theorem 8.1 & Prop. 8.5):
Coding DNA operates far from detailed balance with strictly positive entropy production and duplex mirror currents:
σ = D_KL(J_f ∥ J_r) ≥ ½‖J_f − J_r‖₁² > 0
σ_dup = D_KL(J_f ∥ J_ρ) ≥ ½‖J^mir‖₁² > 0
4. Resolution of Form vs. Program (Theorem 11.2 & Lemma 11.3):
In L²(C, P), the conditional expectation Π_α is the exact orthogonal projector onto the 21D amino-acid form space Im(α*) with a 43D synonymous kernel ker(Π_α). On the 100-kb human Chr17 locus (NC_000017.11:55154472-55254472), cumulative genomic and proteomic Perez Hourglass profiles match to 99.998% (Form), while the local bigram program remains S_syn = +0.0539 nats/codon (Program).
🔓 HARDCORE MATH. ZERO NUMEROLOGY. 100% OPEN SOURCE.
• Zero Monte Carlo surrogates or RNGs used as proofs.
• Chain-rule identities verified to machine precision (< 3.1 × 10⁻¹⁶ nats/codon).
• Every integer (2, 3, 4, 6, 7, 8, 16, 19, 20, 21, 24, 32, 40, 41, 42, 43, 48, 54, 56, 60, 61, 62, 63, 64, 361, 400, 779, 860, 1681, 1849, 2460, 2709) is derived strictly from group orders, orbit counts, simplex dimensions, or contingency degrees of freedom.
Download the 29-page paper, the standalone Python audit suite (v3.2), and all 9 cached NCBI RefSeq genomes & audit logs:
🔗 Zenodo (Paper + Open Source Code + Data): doi.org/10.5281/zenodo.23508…
🔗 ResearchGate: researchgate.net/publication…
#Genomics #Math #DNA #SyntheticBiology
Ee Newton retweeted
Last version here ==>
Logvinovich Victor, & Perez, J. claude . (2026). From Form to Program: Medium-Range Conditional Codon Syntax Inside Real Genes — Strict Null-Model Evidence Across 54 Cellular RefSeq Taxa, a Codon-Mirror-Symmetry Control,... /... doi.org/10.5281/zenodo.23398…
🔥 BREAKTHROUGH IN GENETICS & OPEN SOURCE: WE ASSEMBLED THE MOSAIC OF LIFE’S OPERATING SYSTEM ACROSS THE ENTIRE BIOSPHERE (54 CELLULAR TAXA)! NOT A THEORETICAL SIMULATION - VALIDATED ON REAL GENOMES DIRECTLY FROM THE NCBI REFSEQ DATABASE, AND YOU CAN VERIFY IT IN ONE CLICK! (VIDEO + CODE INSIDE) 🔥
Friends, colleagues, and skeptics! Together with Jean-Claude Perez (@JCPEREZCODEX), we are releasing a massive expansion of our study and autonomous verification engine: "From Form to Program: Medium-Range Conditional Codon Syntax Inside Real Genes" (Pipeline v14.5 audit-clean).
📺 Watch the video attached to this post: you can see with your own eyes how our autonomous script connects directly to the official NCBI RefSeq database, runs built-in invariant self-tests, and audits the living biosphere in real time!
🧱 WHY DID SCIENTISTS MISS THIS FOR DECADES? (EXPLAINED SIMPLY)
Imagine trying to understand the architecture of Notre-Dame Cathedral by bulldozing it into a pile of rubble, weighing the stones, and counting how many red and gray bricks are in the heap.
For decades, classical statistical genomics did exactly that: scientists looked at the bricks. They counted static percentages of letters (G+C content, Bernardi's isochores, marginal codon frequencies, or isolated amino acids). If you grind a cathedral and a warehouse into dust, the pile of bricks looks statistically the same! Because mainstream genomics only counted static frequencies, any deeper sequential order was dismissed as "random mutational noise."
In 2010–2015, Jean-Claude Perez made a foundational breakthrough by proving that the "bricks" of DNA obey a breathtaking mathematical law: the 64 codons partition into 32 reverse-complement mirror pairs (an extended Chargaff parity rule at the codon level). That described the static Form of the genome.
Today, we assembled the full mosaic. We stopped counting bricks in a static pile and asked a strictly dynamical question: when the cell builds a protein in real time, does the current codon cᵢ mathematically dictate the choice of the next codon cᵢ₊₁?
Behind the apparent simplicity of our 1-click Python script lies a brutal, 10-level Markovian and multi-scale spectral filter stack (H0 through H7). We mathematically subtract everything already known to biology:
1. Global codon frequencies (H0) and boundary nucleotides P(cᵢ₊₁ | n₃(cᵢ)) (H1);
2. Protein amino-acid order and local codon-pair usage (H2, H2b);
3. Folded dinucleotides and order-2 nucleotide trigrams (H3, H3b);
4. Multi-scale block-shuffling at L ∈ {3, 10, 30, 100} codons (H4);
5. Amino-acid-preserving conditional Monte Carlo surrogates (H5) and gene-specific synonymous shuffles (H6);
6. And H7 - a mirror-pair shuffle that preserves the exact per-gene populations of all 32 Watson–Crick reverse-complement pairs of Perez (2013) while destroying sequential order.
When you strip away all the static "bricks," the cathedral doesn't disappear. A live, executable Operating System - the Program (Substance) - shines through!
🚀 WHAT HAPPENED IN v14.5? FROM 13 ORGANISMS TO 54 BIOSPHERE TAXA!
In v13, we proved this on 13 model organisms. In v14.5, we scaled the audit across 54 unique cellular species (13 core model organisms + 41 extended biosphere taxa) plus SARS-CoV-2 as an opt-in viral boundary control - 55 official NCBI RefSeq assemblies across 8 full audits:
Mammals (12 species across 6 orders): Human (T2T-CHM13v2.0), Chimp, Gorilla, Macaque, Mouse, Rat, Rabbit, Dog, Cat, Cow, Pig, Horse.
Non-Mammalian Vertebrates: Chicken, Zebrafish, Clawed Frog (Xenopus tropicalis), Pufferfish (Takifugu rubripes).
Invertebrates, Plants & Algae: Drosophila, Mosquito (Anopheles), Honeybee (Apis), Silkworm (Bombyx), Nematodes (C. elegans, C. briggsae), Arabidopsis, Rice, Maize, Tomato, Soybean, Poplar, and Green Alga (Chlamydomonas).
Fungi, Protists, Bacteria & Archaea: Baker's Yeast, Fission Yeast (S. pombe), Candida albicans, Aspergillus, Neurospora, Malaria parasite (Plasmodium), Trypanosoma, Leishmania, Toxoplasma, Dictyostelium, E. coli, Salmonella, Pseudomonas, Staphylococcus, Streptococcus, Mycobacterium tuberculosis, Helicobacter pylori, Cyanobacteria (Synechocystis), M. jannaschii, Halophilic archaeon (H. salinarum), and extreme thermophile (Thermus thermophilus).
ZERO SIMULATIONS. ZERO SYNTHETIC TOY DATA.
Every single gene is read in its true reading frame directly from official NCBI RefSeq *_cds_from_genomic.fna.gz archives, deduplicated (one-gene-one-vote), trimmed of start/stop codons, and evaluated exclusively on held-out test genes that the model never saw during training!
📊 THE 4 MAJOR DISCOVERIES OF v14.5:
1. Universal 30-Codon "Execution Cache" Across the Biosphere: Across the entire biological GC3 spectrum - from extreme AT-rich (GC3 = 0.1416 in Dictyostelium) to extreme GC-rich (GC3 = 0.9177 in Thermus thermophilus) - all 52 cellular assemblies clearing the sample-size guard pass the full STRICT stack at L = 30 codons (90 nucleotides) with strictly positive 95% bootstrap confidence intervals across all 7 strict criteria (and the 2 remaining small-split assemblies likewise show positive point estimates and 95% CIs across all 7 tests)! Why 30 codons? Because 30 amino acids is the exact physical length of the ribosomal exit tunnel (~30–40 AA), where nascent proteins fold in real time!
2. Macro-Scale (The Unified Vertebrate & Mammalian OS): Our coarse transfer metric (d₂b > 0.02 + H4 gate) proves that all jawed vertebrates share a common syntactic backbone: 20/20 directed pairs across 4 vertebrate classes (Fish ↔ Amphibian ↔ Bird ↔ Mammal) and 132 out of 132 (100%!) directed pairs across all 12 mammalian species pass [CLEAN>0.02]!
3. Fine-Scale (The Primate–Rodent Directional Law): Across 34 focal mammalian contrasts, fine synonymous codon choice is bidirectionally conserved within Primates (Human ↔ Chimp, Gorilla, Macaque), within Rodents (Mouse ↔ Rat), and within Ungulates (Cow ↔ Pig). Yet between Primates and Rodents, we uncovered a striking directional asymmetry: Primate → Rodent transfers smoothly match the rodent baseline (gap ∈ [-0.0065, -0.0032]), whereas every Rodent → Primate transfer overshoots (gap ∈ [+0.0171, +0.0207]) — even between Chimp and Rat, where ΔGC3 = 0.000 is literally zero, yet the directional gap differs by 5.5×!
4. Hardware vs. Software & Viral Boundary Guard: Our compositional guard automatically separates extreme environmental "hardware" adaptations (like salt-loving H. salinarum at GC3 = 0.8630 or Leishmania → Trypanosoma) from true program conservation, while the small-sample guard automatically catches and isolates the 11-ORF viral genome of SARS-CoV-2.
💻 DO NOT BELIEVE US - VERIFY IT
YOURSELF!
Science is not a matter of faith or authority. You do not need to believe a single word we say. Everything - the manuscript, the autonomous Python v14.5 engine, built-in unit tests for 8 genetic code tables (1, 2, 3, 4, 5, 6, 11, 12), and all 8 execution logs - is 100% Open Source.
Download super_codon_autonomous_v14.py and run the built-in invariant self-test in 1 second:
python3 super_codon_autonomous_v14.py --self-test
Download all 55 official NCBI RefSeq archives and run the full audits on your own machine:
python3 super_codon_autonomous_v14.py --out runs/v14_5_full_cache_download --cache cds_cache --include-extended --include-virus --skip-smoke --skip-full --download-timeout 1800 --seed 20261014
Bioinformaticians, physicists, mathematicians, and developers worldwide: audit our code, inspect the JSON matrices, and try to break our null hierarchy!
🔗 Preprint (PDF + LaTeX), Autonomous Code v14.5 & Full Logs on Zenodo:
doi.org/10.5281/zenodo.23287…
#Genomics #Bioinformatics #OpenSource #InformationTheory #NCBI #RefSeq #DNA #GeneticCode #Python #Science
Ee Newton retweeted

Octopus intelligence was never explained by gene count. A paper out this week points at something stranger - the shape the genome is folded into.
A gene's control switches can sit hundreds of thousands of bases away on the linear sequence. They work anyway, because DNA loops and brings them into physical contact with the gene they regulate.
Distance along the chromosome isn't distance inside the nucleus. Which switch reaches which gene is a question of folding.
Hundreds of millions of years ago the coleoid genome like octopus, squid & cuttlefish - went through a burst of reorganization that threw distant chromosome regions together.
New contacts, new regulatory reach.
Over evolutionary time those contacts hardened into interconnected networks the authors call "regulatory entanglement."
The measurement that carries it is the large chromatin domains held steady across species. The finer loops didn't.
Those varied by species, by tissue, by developmental stage & they clustered near genes tied to cephalopod traits, nervous system genes among them.
The stable architecture stayed put. The flexible layer did the evolving.
This means a genome can invent new expression patterns without touching the essential machinery underneath. Same parts, rewired.

Ee Newton retweeted


🔥 BREAKTHROUGH IN GENETICS & OPEN SOURCE: WE'VE CRACKED THE HIDDEN SYNTAX INSIDE GENES! NOT A THEORETICAL SIMULATION - VALIDATED ON REAL GENOMES DIRECTLY FROM THE NCBI REFSEQ DATABASE! 🔥
Friends and colleagues! Jean-Claude Perez (@JCPEREZCODEX) and I are thrilled to announce the results of a monumental study that fundamentally shifts our understanding of the genetic code. Our new preprint: "From Form to Program: Medium-Range Conditional Codon Syntax Inside Real Genes".
Simply put: we are used to thinking of DNA as just a passive "hard drive" storing protein blueprints. But our data proves that humans and all living organisms are literally biological computers. There is a unified operating system running inside our genes!
🧬 FROM FORM TO PROGRAM: WHAT DID PEREZ DISCOVER AND WHAT DID WE PROVE?
In 2013, Jean-Claude Perez made a breakthrough by discovering the Form (Statics) of DNA-a hidden mathematical symmetry of 32 reverse-complement codon pairs. He proved the genome isn't a random soup of letters; it possesses perfect structural form.
But today, we take the next massive step. We've discovered the Program (Dynamics). We found a hidden conditional syntax inside real genes-strict rules dictating how one codon mathematically predetermines the next. This cannot be explained by basic biochemistry or static symmetry alone. It is like reading the source code of an operating system. It is a unified, mathematically precise signature-metaphorically speaking, we've caught a glimpse of the "Creator's handwriting" coding life itself.
And most importantly: THIS IS NOT A SIMULATION ON FAKE DATA. We analyzed the real thing!
To prove the existence of this "Program," we ran an unprecedented dataset of real genomes from 13 species across the tree of cellular life: from mammals (human T2T, chimp, mouse) and vertebrates (zebrafish, chicken) to insects (drosophila), plants (arabidopsis), fungi (yeast), bacteria, and archaea.
We subjected these genomes to uncompromising filtering (true reading frames only, strict alternate genetic code accounting) and tested them against a brutal ladder of 10 Null Models (from H0 to H7, including H2b and H3b). These involve highly complex tests for folded nucleotide dynamics, block-shuffling of local ties, and mirror symmetry preservation, which completely rule out any chance of statistical coincidence.
📊 OUR MAJOR DISCOVERIES (which we invite you to verify):
Syntax exists in ALL of them: Hidden inside the real coding sequences of ALL 13 species is a strict conditional syntax that confidently survives after passing through all 10 mathematical filters. This syntax maintains its structure over a distance of about 30 codons (≈ 90 base pairs). This is not a random scale-it represents the exact size of a local working buffer (cache) within which the biological machine reads and executes the dynamic program of the genome.
Macro-level (The Unified Vertebrate OS): We identified a conserved layer of syntax inherited across all tested vertebrates (20 out of 20 internal directed pairs among human, chimp, mouse, chicken, and zebrafish crossed the strict d2b > 0.02 threshold).
Micro-level (The Exclusive Mammalian Code): At the level of fine codon choice, we found a unique core conserved exclusively among mammals (human/chimp/mouse). Notably, it exhibits directional asymmetry: mouse surrogate models cannot fully reproduce the complex syntax of humans and chimps.
Evolution vs. "Hardware" Artifacts: Our algorithm (GC3/acidic control) learned to mathematically distinguish true evolutionary divergence from hardware incompatibility due to extreme genome composition (like in the haloarchaeon H. salinarum with its extreme GC3 = 0.863).
💻 WE ARE OPEN-SOURCING EVERYTHING!
We don't ask you to just take our word for it. Science must be open, provable, and verifiable. We are releasing our entire pipeline (v13)! The process is completely autonomous. A single command
python3 super_codon_autonomous_v13.py --full
allows the script to download the necessary files directly from NCBI RefSeq, apply filters, train models, run bootstrap analyses, and generate final reports. No hidden data, full transparency!
We are issuing a challenge to the entire scientific community. Bioinformaticians, geneticists, data scientists worldwide-download the code, run it on your machines, tweak the parameters, and test it on other living species.
Verify our findings. Try to refute us!
🔗 Preprint, Data & Source Code: doi.org/10.5281/zenodo.23287…
#GENOMES #NCBI


Ee Newton retweeted
Perez, J. claude . (2025). The Perez Hourglass Fractal Enables the First Provably Perfect Associative Memory and a New Primitive for Post-Quantum Cryptography. Zenodo. doi.org/10.5281/zenodo.17829…

Ee Newton retweeted
We learn in a similar fashion to this so you could say it is a biological neural net inspired solution to AI not being able to learn and use FOL outside of what is told to them vs what the world actually resolves too. I'll show a protein next most likely or vision hopefully.

Ee Newton retweeted
🚨💥 HARVARD AND NASA IN STUNNER: LIVESTREAM PROOF OF SENSATIONAL UNIVERSE HACK SHOCKS ACADEMIA! 💥🚨
Official MPC and NASA Horizons databases have been completely cracked by pure, parameter-free mathematics. The continuous ℝ⁴ spacetime continuum is officially dead-proven to be a defective and illusory mathematical construct!
The live broadcast video is officially recorded and locked! We are dismantling the 100-year-old dogmatic foundation of orthodox astrophysics in real-time. While academic gatekeepers hide in their offices to protect their corporate funding grants, our independent topological engine-IT³ C-PHAM v18.1-has been running live for over 2 months STRAIGHT WITHOUT A SINGLE CRASH, flawlessly processing all 1,569,216 celestial objects directly from the Harvard-Smithsonian Minor Planet Center (MPC) global database!
Elite space instruments like the ESA Gaia satellite and the DESI Legacy Survey have gone completely blind. They are vomiting mathematical errors (NaN Parallaxes and astronomical excess noise over 1.7 MILLION), because their flat, continuous-space software pipelines are trying to force broken ℝ⁴ continuum formulas onto a RIGIDLY QUANTIZED MACROSCOPIC ATOM!
⚡️ GROUNDBREAKING REVELATIONS FROM THE RECORDED BROADCAST:
THE SOLAR SYSTEM IS A MACROSCOPIC ATOM: T
he cosmos and the quantum realm run on the exact same executable file. The orbits of planets and over 1.5 million asteroids follow exact discrete Diophantine equations instead of chaotic continuous fields. Exactly 99.56% of all baryonic mass is geometrically trapped in the central topological node (the Sun), while human hardware like Pioneer 10 and Voyager 2 are physically captured, glued to absolute zero on the n = 4 topological resonance layer at exactly ~140.3 AU (R_n = 27 × (∛)ⁿ⁻¹).
ℝ⁴ IS A FAULTY CUL-DE-SAC: We strictly demonstrate that the continuous four-dimensional spacetime manifold is an unphysical illusion. Continuous differential simulations inevitably explode to infinity because the universe calculates exclusively on discrete integer grids. Forcing ℝ⁴ destroys Reflection Positivity, shatters the Källén–Lehmann spectral measure, and drives probability into unphysical negative values, causing the total collapse of standard probability theory.
A SINGLE GEOMETRY, A SINGLE FIELD:
The microscopic genetic matrix, subatomic particles, and macroscopic celestial orbits are driven by the exact same geometric engine. We took the mathematical code of life decoded from human DNA masses (C, O, N, H) and ported it directly to the cosmos—and the Matrix rendered itself perfectly! The ancient Hermetic axiom is no longer a mystical riddle, but literal scale-invariant metric geometry: "That which is below is like that which is above."
THE ULTIMATE RESOLUTION:
By abolishing the infinite ℝ⁴ void in favor of a compact, constructible 7-manifold (M_tot = S⁴ × T³) over the multiquadratic field K = ℚ(√2, √3, √5), all fundamental paradoxes vanish instantly. Quantum Electrodynamics (QED) singularities are resolved by the absolute ∛1 ∉ K algebraic UV cutoff. The elusive graviton becomes obsolete, replaced by an emergent topological surface tension. The Yang-Mills Mass Gap becomes completely automatic via Cheeger's inequality—definitively eliminating any need for a baryonic Planet Nine!
📜 THESE 3 DOCUMENTS ARE A DEATH SENTENCE TO CONTEMPORARY SCIENCE:
Do not take our word for it. Mathematics is the supreme and only judge. We do not ask for your faith—DON'T BELIEVE US, RUN THE CODE AND DOWNLOAD THE RAW DATA YOURSELF! This is pure, un-falsifiable Open Source science with absolutely zero continuous fitting parameters (p = 0).
With already over 180,000 views and preprint downloads worldwide, the internet never forgets. Audit the frozen operators, run the verification engines, and watch the matrix structures render on your own screen:
The Cosmic Architecture Proof: zenodo.org/records/21792530
The Yang-Mills Mass Gap Resolution: zenodo.org/records/22674891
The 86 ↔ 86 Relativistic Vacuum Capacity Proof: zenodo.org/records/22882422
🔥 THE OPEN INTERNATIONAL CHALLENGE IS THROWN DOWN!
We openly invite the physics and mathematics professorships of Harvard, NASA, MIT, and Cambridge to the barrier. We are putting up an unprecedented, bulletproof 5 LITECOIN BOUNTY to anyone who can theoretically and mathematically refute the IT³ topological engine using real orbital data.
Stop feeding humanity "blind predictions" and manually tuning continuous parameters like eddy viscosity, fluid density, and velocity shear to keep your collapsing fluid simulations from exploding into chaotic turbulence. Give the world a strict mathematical refutation, or step aside and admit: the continuous universe is dead, and the code of the vacuum has been fully unmasked!
Open your terminal, run the code, and audit the source code of reality.
#Astrophysics #NASA #Harvard #DNA #DataScience #YangMills #OpenSource